Injectable Packaging Integrity Testing: Explore How Seal Integrity Works
Injectable medicines depend on packaging that can maintain product protection from filling through storage and administration.
Injectable packaging integrity testing evaluates whether the container and closure system can prevent unwanted leakage, contamination, or loss of protection. For sterile products, even a small defect can have consequences for product quality.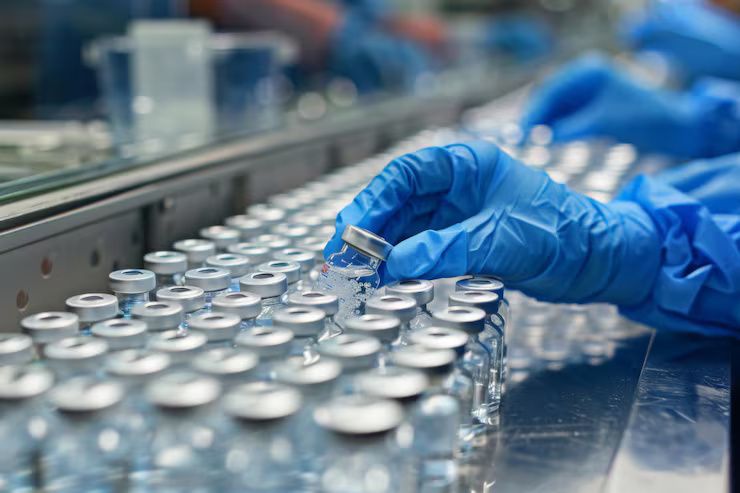
Modern injectable packaging may include glass vials, prefilled syringes, ampoules, cartridges, elastomeric stoppers, caps, seals, and administration components. Each interface can influence package integrity, making the relationship between the container, closure, sealing process, and product environment important during development and manufacturing. FDA guidance describes container and closure integrity as part of maintaining sterility throughout a sterile product's shelf life.
Understanding how seal integrity works helps explain why packaging is tested under controlled conditions rather than judged only by visual appearance. It also clarifies the difference between detecting a physical leak, evaluating seal performance, and demonstrating that a package can maintain its intended protective function over time.
What Seal Integrity Means in Injectable Packaging
Seal integrity refers to the ability of the assembled container closure system to maintain an effective barrier between the drug product and its surrounding environment. The assessment concerns the complete packaging system rather than one component in isolation.
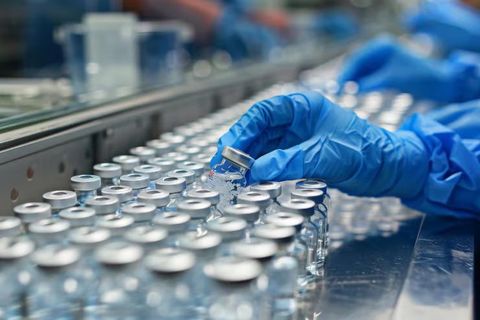
For a vial, for example, the system can include the glass container, elastomeric stopper, aluminum overseal, and the way those components interact after stoppering and crimping. A prefilled syringe introduces different interfaces involving the barrel, plunger stopper, tip closure, and other components.
A package may appear intact while still containing a microscopic pathway that permits gas, moisture, microorganisms, or liquid to move through the closure system. This is why container closure integrity testing uses controlled physical or chemical detection methods capable of identifying leakage that visual inspection cannot reliably establish. FDA identifies methods such as pressure or vacuum decay, tracer-gas testing, dye penetration, and other validated approaches as possible integrity-testing techniques.
The central principle is straightforward: the package must remain sufficiently closed to protect the product under its intended conditions of storage and use.
How the Seal Actually Creates a Barrier
Seal performance begins with the interaction between packaging materials. In many injectable systems, an elastomeric closure is compressed against a rigid container surface. The resulting contact area forms a barrier that limits pathways through which contaminants or product could pass.
The effectiveness of that barrier depends on factors such as compression, component dimensions, material properties, surface condition, assembly forces, and manufacturing tolerances. A closure that is physically present is not necessarily a closure that has formed an adequate seal.
The sealing process also matters. During vial packaging, the stopper must be correctly positioned before the overseal is applied. Crimping then helps maintain the closure configuration. Variations in component dimensions, equipment settings, stopper placement, or crimping conditions can influence the final package.
For this reason, integrity testing is closely connected with packaging design and process validation. FDA guidance recommends that integrity methods be appropriate for the particular product and container closure system and be validated for their intended purpose.
Common Ways Injectable Package Integrity Is Tested
No single testing method is suitable for every injectable package. The method must match the package design, expected leak characteristics, required sensitivity, and intended application.
Vacuum and Pressure Decay Testing
Vacuum decay testing places a package inside a controlled chamber and creates a vacuum around it. A leak can cause pressure changes within the chamber, which are detected by pressure sensors.
ASTM F2338 describes nondestructive leak detection using vacuum decay. The technique measures changes associated with leakage from the package or, in some circumstances, volatilization of liquid near a leak.
Pressure decay follows a related principle but evaluates pressure changes under a controlled pressurized condition. These approaches can be useful when nondestructive testing is desirable and the package configuration is compatible with the equipment.
Helium Tracer-Gas Testing
Helium testing uses helium as a tracer gas because its behavior allows highly sensitive leak detection. Depending on the procedure, helium may be introduced into the package or incorporated into the headspace before sealing.
The sealed package is then evaluated using a helium leak detector. ASTM F2391 describes both vacuum-mode and sniffer-mode approaches and recognizes applications ranging from packaging development to process validation and quality monitoring.
Helium testing can provide high sensitivity, but the selected procedure must account for package design, background helium, fixtures, permeability, and other sources of measurement variability.
Dye Penetration and Visual Methods
Dye-based methods can help identify leakage pathways by allowing a detectable dye to penetrate through defects under controlled conditions. These methods can be useful for certain packaging configurations, particularly where the location or presence of a gross defect needs to be demonstrated.
Visual inspection remains useful for identifying obvious defects such as damaged components, improper assembly, or visible seal abnormalities. However, visual inspection alone cannot establish that a package is microbiologically or physically integral at the required sensitivity.
The appropriate method therefore depends on what the test needs to demonstrate rather than on selecting the most sophisticated technique available.
From Leak Detection to Seal Validation
A major distinction in injectable packaging is the difference between performing a leak test and validating that the packaging process consistently produces an acceptable seal.
Validation typically considers the entire system. Engineers may evaluate representative packaging configurations, deliberately introduced defects, process variables, environmental conditions, and the sensitivity of the selected test method.
Acceptance criteria should be connected to the intended package performance. A generic statement such as “no leaks detected” is not necessarily sufficient unless the test method has appropriate sensitivity and the detection threshold is scientifically justified.
FDA guidance emphasizes that container closure integrity methods should be validated and capable of detecting a breach relevant to the specific container and closure system.
This makes method sensitivity particularly important. A test that cannot reliably detect the relevant defect size may produce a passing result without providing meaningful assurance about package integrity.
Why Manufacturing Conditions Matter
A validated package design can still encounter integrity problems if the manufacturing process is poorly controlled. Component variability, equipment alignment, sealing pressure, temperature, handling, and assembly conditions can all affect the final closure.
For injectable products, this is particularly significant because packaging must maintain its protective function through transportation and storage, not merely immediately after sealing.
Packaging studies may therefore evaluate samples from different production conditions and stages. Testing can also be incorporated into development, equipment qualification, process validation, stability programs, and ongoing quality activities.
The objective is not simply to identify defective packages after production. A stronger approach uses testing data to understand which process variables influence seal performance and how those variables can be controlled.
Integrity During Storage and Use
Package integrity is not necessarily a static property. Temperature changes, mechanical stresses, material aging, pressure differences, and interactions between the drug product and packaging components can influence performance over time.
For sterile products, maintaining integrity throughout the labeled shelf life is particularly important. FDA explains that container closure integrity testing can be used within appropriate stability programs to help demonstrate continued protection of sterile products, although such testing does not replace release sterility testing.
Injectable containers can also have specific in-use requirements. For example, multidose containers may need to maintain closure functionality after repeated needle penetrations. FDA notes that functionality testing for multidose injection containers can include evaluation of self-sealing capacity after multiple penetrations.
This highlights an important point: package integrity is not limited to the moment a container leaves the filling line. The packaging must perform according to its intended use throughout the product lifecycle.
Selecting an Appropriate Testing Strategy
An effective injectable packaging integrity program starts with understanding the package rather than selecting a test method first. Important considerations include the container material, closure design, sealing mechanism, product characteristics, intended storage conditions, and how the package will be accessed.
The testing strategy should also distinguish between development testing, process validation, stability testing, and routine quality monitoring. Different stages may require different methods or sampling approaches.
Regulatory expectations also favor scientifically justified methods. FDA states that testing should be appropriate to the specific container closure system and product, with validation demonstrating that the method can detect relevant breaches.
In practice, this means that a robust program combines packaging engineering, analytical testing, manufacturing controls, and documented acceptance criteria rather than relying on one inspection technique.
Frequently Asked Questions
Does a visual inspection prove injectable packaging is sealed?
No. Visual inspection can identify obvious defects, but it generally cannot establish the absence of small leaks or demonstrate complete container closure integrity.
Is container closure integrity testing the same as sterility testing?
No. Integrity testing evaluates whether the packaging can maintain its protective barrier. FDA specifically distinguishes container closure integrity testing from sterility testing performed for product release.
Which test method is appropriate for every injectable package?
There is no universal method. The selected technique should match the package design, required sensitivity, product characteristics, and intended use.
Why is seal validation performed beyond initial packaging?
Because package performance can change during storage, transportation, environmental exposure, and intended use. Stability-related integrity testing can help demonstrate continued protection over the product's dating period.
Conclusion
Injectable packaging integrity testing provides a structured way to determine whether a container closure system can maintain its intended protective barrier. Seal integrity depends on material interaction, closure design, assembly conditions, manufacturing controls, and performance over time.
Vacuum decay, pressure-based methods, helium tracer-gas testing, dye techniques, and other approaches can each have appropriate applications. The critical factor is not choosing the most advanced test, but selecting and validating a method that can reliably detect the type of package defect relevant to the injectable product.
Related Post...
Biotech Pipeline: Explore Trends Across Clinical Development Stages
Updated: August 22, 2026
Read More
Dental Clips in Orthodontics Discover Modern Teeth Alignment Solutions
Updated: July 11, 2026
Read More